
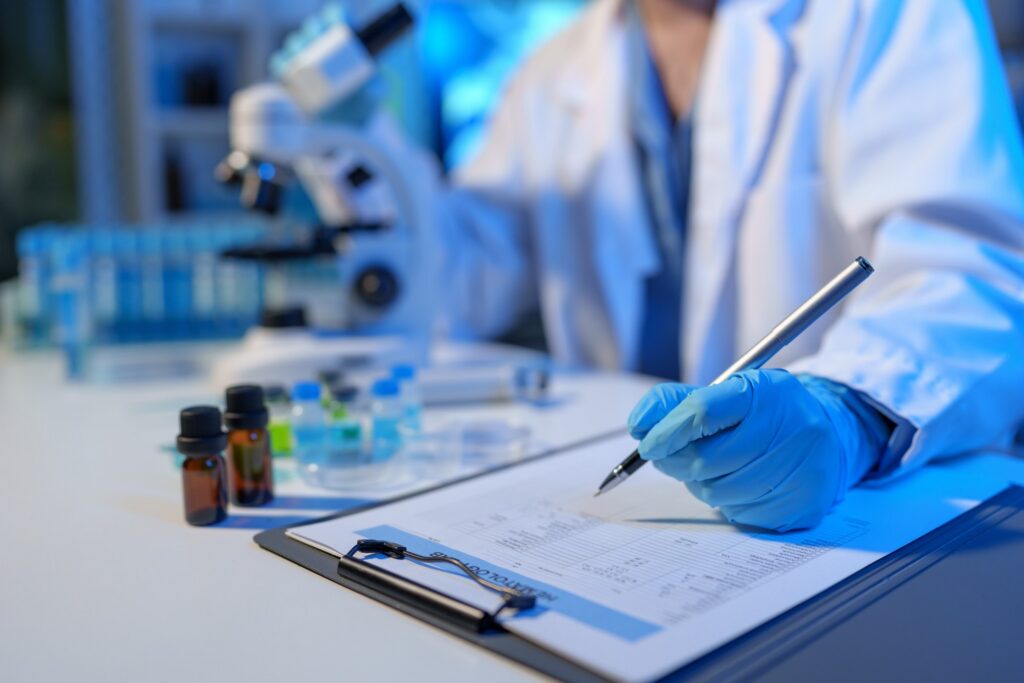
Regulatory affairs for diagnostics, medical devices and digital health
Abingdon AI & MedTech
Regulatory affairs
Bringing a medical device, diagnostic or digital health technology to market involves far more than just submitting documentation. Regulatory requirements influence development decisions, evidence generation, quality systems and commercial timelines. Abingdon AI & MedTech helps organisations build a clear route to market strategy and navigate regulatory milestones across the US, EU, UK and over 90 international markets.
What we deliver
Regulatory affairs services
MedTech Regulatory Inspection Gap Analysis
Regulatory inspections rarely fail because of a single issue. More often, findings reveal underlying weaknesses in documentation, procedures or quality oversight that have developed over time. Our MedTech regulatory Gap Analysis service identifies where those vulnerabilities exist, benchmarks them against current FDA, ISO 13485, EU/UK MDR and IVDR expectations, and provides a practical roadmap for remediation and inspection readiness.
FDA Submissions (510(k), PMA, De Novo)
FDA submissions are rarely just administrative exercises. The strongest medical device regulatory programmes are built around a clear understanding of regulatory expectations long before a submission is drafted. We support organisations throughout the process, from product qualification, product classification, regulatory pathway assessment, predicate device evaluation and Q-Submissions (pre-submissions) through to 510(k), De Novo and PMA submission preparation, FDA interactions and review support.
CE Marking & UKCA
Obtaining CE Marking or UKCA certification is often straightforward in principle but challenging in practice. Questions around medical device classification for regulatory approval, evidence requirements and notified body expectations can quickly delay progress. We help manufacturers establish clear regulatory pathways and prepare the documentation needed to move products towards approval, drawing on proven expertise.
International Regulatory Submissions
Expanding medical device sales into international markets requires coordinated regulatory planning across multiple territories. We support submissions and registrations across key global markets, including the US, EU, UK, Australia, Brazil and APAC regions, providing comprehensive support for FDA, CE, UKCA and over 90 territories globally.
MDR & IVDR Consultation
Many manufacturers continue to face uncertainty under the Medical Devices Regulations (UK MDR 2002), EU MDR 2017/745 and the EU In Vitro Diagnostic Regulation 2017/746 (IVDR), whether developing a new device for market, transitioning legacy products, addressing documentation deficiencies or responding to notified body feedback. Our team provides hands-on support to help organisations maintain compliance, protect market access and navigate evolving market requirements under these regulations.
Product Qualification & Classification
Product qualification and classification influence almost every subsequent regulatory decision, from evidence requirements and conformity assessment routes through to timelines and costs. Getting it wrong early can create significant delays later. We help manufacturers establish the appropriate classification and understand the implications for development, approval and commercialisation.
Medical Device Post-Market Surveillance, Vigilance & Reporting
Maintaining compliance after market launch requires effective vigilance and Post-Market Surveillance (PMS) systems. We support complaint handling, Medical Device Reporting, vigilance reporting, recalls, PMS, PMCF, PMPF and corrective action activities, helping manufacturers meet regulatory obligations while protecting patient safety and maintaining market access.
Our approach
Bringing an AI or Digital Medical Device to Market?
Abingdon AI & MedTech combines regulatory, quality and clinical expertise with broader capabilities across product development, analytical services and commercialisation. Our team supports developers and manufacturers of molecular diagnostics, immunoassays, point-of-care testing, lateral flow tests, in vitro diagnostics, medical devices, healthcare software, SaMD, digital health and AI-enabled technologies across the complete product lifecycle.
From the Knowledge Centre
Read more on Regulatory affairs
Practical guidance from the specialists who deliver this work every day.
 Article
Performance Evaluation for FDA 510(k): Building Your Substantial Equivalence Case
What FDA 510(k) performance evaluation requires and how to build a strong case for substantial equivalence in IVD submissions.
Also worth reading
-
 Article
Companion Diagnostics: Playing 5D Chess in Regulatory Development
-
 Article
Same Innovation, Different Pathways: AI Regulation in the US, EU & UK
-
 Article
Why ISO/IEC 17025 Accreditation Matters to Your Regulatory Submission
-
 Article
Navigating new UK medical device regulations: Action required now, help is on hand!
Case study
Find Out From Home STD Tests Development Project
How Abingdon Health partners with Find Out From Home (FOFH) to develop and commercialise rapid sexually transmitted disease self-tests.
Frequently asked questions
Regulatory affairs FAQs
We provide regulatory strategy, product classification, FDA submissions, CE marking, UKCA support, MDR and IVDR consultation, international registrations and post-market compliance services. Support is available across medical devices, in vitro diagnostics, Software as a Medical Device (SaMD) and digital health technologies.
The appropriate regulatory pathway depends on the product’s intended use, classification, technology and target market. Early product qualification and classification can help identify the evidence, documentation and regulatory requirements needed for approval.
Support includes regulatory pathway assessment, product classification, predicate device evaluation, pre-submissions, submission preparation and regulatory interactions. A structured FDA strategy can help reduce delays and strengthen submission readiness.
MDR applies to medical devices, while IVDR applies to in vitro diagnostic medical devices. Each regulation has different classification rules, evidence requirements and conformity assessment processes that manufacturers must address before placing products on the market.
Yes. We support CE marking, UKCA certification, international registrations and regulatory submissions across the United States, European Union, United Kingdom and more than 90 international markets.
Manufacturers are required to maintain post-market surveillance, vigilance reporting and ongoing compliance activities following product approval. These activities may include complaint handling, PMS, PMCF, PMPF, corrective actions and regulatory reporting obligations.
Start the conversation
Discuss your regulatory strategy with us
Whether you are preparing for FDA submission, pursuing CE Marking or UKCA certification, transitioning to MDR or IVDR, or planning global market expansion, Abingdon AI & MedTech can support your programme. Speak with our experienced team of regulatory professionals to discuss your product, regulatory challenges and route to market.
- US, EU and UK market access
- ISO 13485, MDR, IVDR and FDA expertise
- Flexible support models